AlignBench: pairwise DNA alignment

AlignBench compares two DNA sequences using the same algorithms and default scores as EMBOSS needle and water, and shows the alignment with mismatches and gaps highlighted.

Open AlignBenchAll tools

Free, no sign-up, and installable on your phone. Runs in your browser at align.gataca.com.
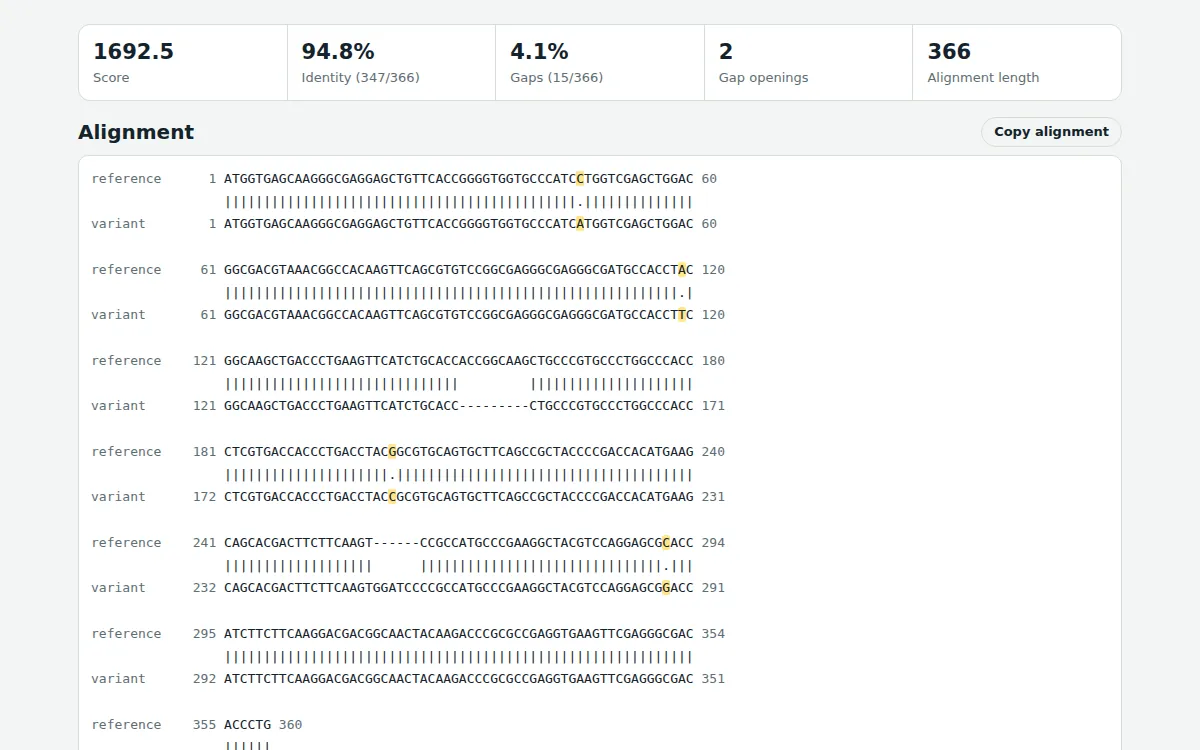
AlignBench showing results for the built-in example
AlignBench with its built-in example loaded.

What you can do with AlignBench

  • Confirm a Sanger sequencing read against your reference
  • Spot point mutations, insertions, and deletions in a clone
  • Find where a short fragment sits within a longer sequence
  • Compare homologous genes or alleles

How to use it

  1. Paste the reference as sequence A and the read or variant as sequence B.
  2. Choose Global to compare end to end, or Local to find the best-matching region.
  3. Adjust match, mismatch, and gap scores if needed, or keep the EMBOSS defaults.
  4. Review identity, gaps, and the highlighted alignment, then copy it.

Features

  • Global alignment (Needleman-Wunsch)
  • Local alignment (Smith-Waterman)
  • Affine gap penalties (Gotoh algorithm)
  • EMBOSS-compatible default scoring
  • Reverse complement option for sequence B
  • Identity, gap, and score statistics
  • Save results as reports, spreadsheets, or sequence files
  • Install it on your phone or computer; works offline

Frequently asked questions

Why use affine gap penalties?

Real indels usually span several bases. Charging more to open a gap than to extend it produces alignments that match biology better.

My read aligned poorly. What should I try?

If the read is from the opposite strand, tick the reverse complement option. If it covers only part of the reference, switch to Local.

Can AlignBench align proteins or many sequences?

It currently aligns two DNA sequences. For multiple sequence alignment, tools such as Clustal Omega or MAFFT are the right choice.

More free tools

Need AlignBench to do more?

We build custom versions and full pipelines for research teams.
Talk to us